
International Drug Regulatory Affairs 1 Regulatory


Description: International Drug Regulatory Affairs 1 Regulatory structure The pharmaceutical industry is one of the highly regulated industries with various structures of drug regulation Drug laws Drug regulatory agencies Drug evaluation boards
Related Topics
Download Presentation
"International Drug Regulatory Affairs 1 Regulatory" is the property of its rightful owner. Permission is granted to download and print the materials on this website for personal, non-commercial use only, and to display it on your personal computer provided you do not modify the materials and that you retain all copyright notices contained in the materials. By downloading content from our website, you accept the terms of this agreement.
Presentation Transcript
slide1. International
Drug Regulatory Affairs 1<br>
slide2. Regulatory structure The pharmaceutical industry is one of the highly regulated industries with various structures of drug regulation
▫ Drug laws
▫ Drug regulatory agencies
▫ Drug evaluation boards
▫ Quality control laboratories
▫ Drug information centers 2<br>
slide3. What is the need? Drug laws provide the basis for drug regulation
Regulatory tools such as standards and guidelines equip authorities with the practical means of implementing those laws
Regulatory Agencies monitor the implementation of these laws 3<br>
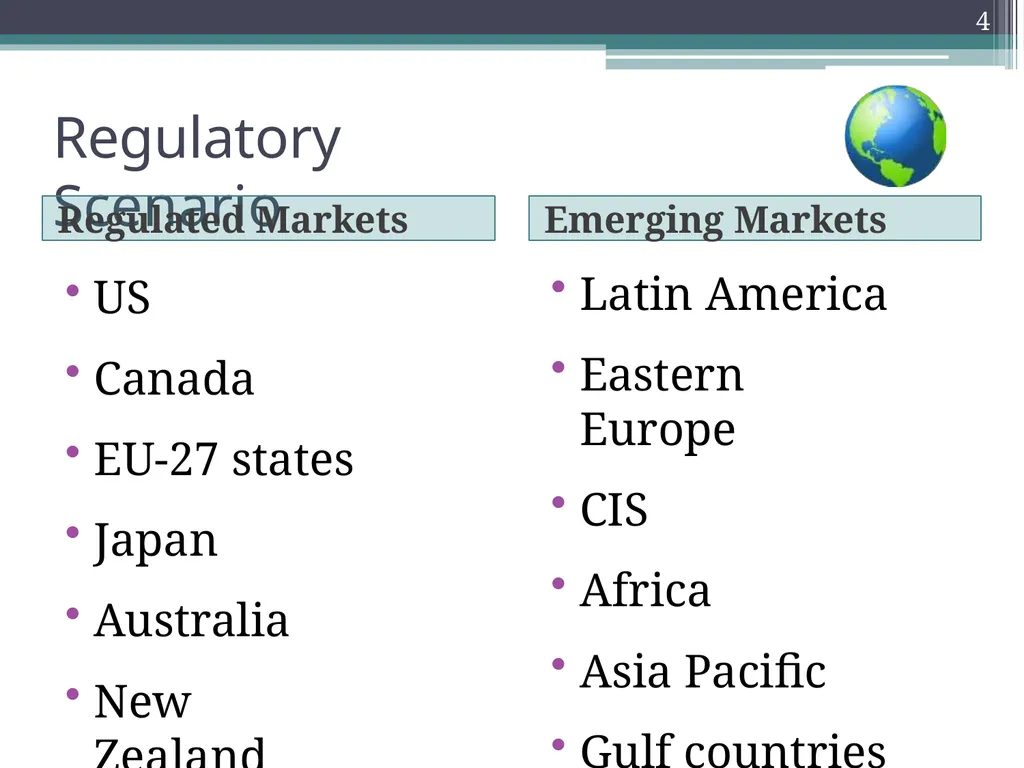
slide4. Regulatory Scenario Regulated Markets Emerging Markets US
Canada
EU-27 states
Japan
Australia
New Zealand Latin America
Eastern Europe
CIS
Africa
Asia Pacific
Gulf countries 4<br>
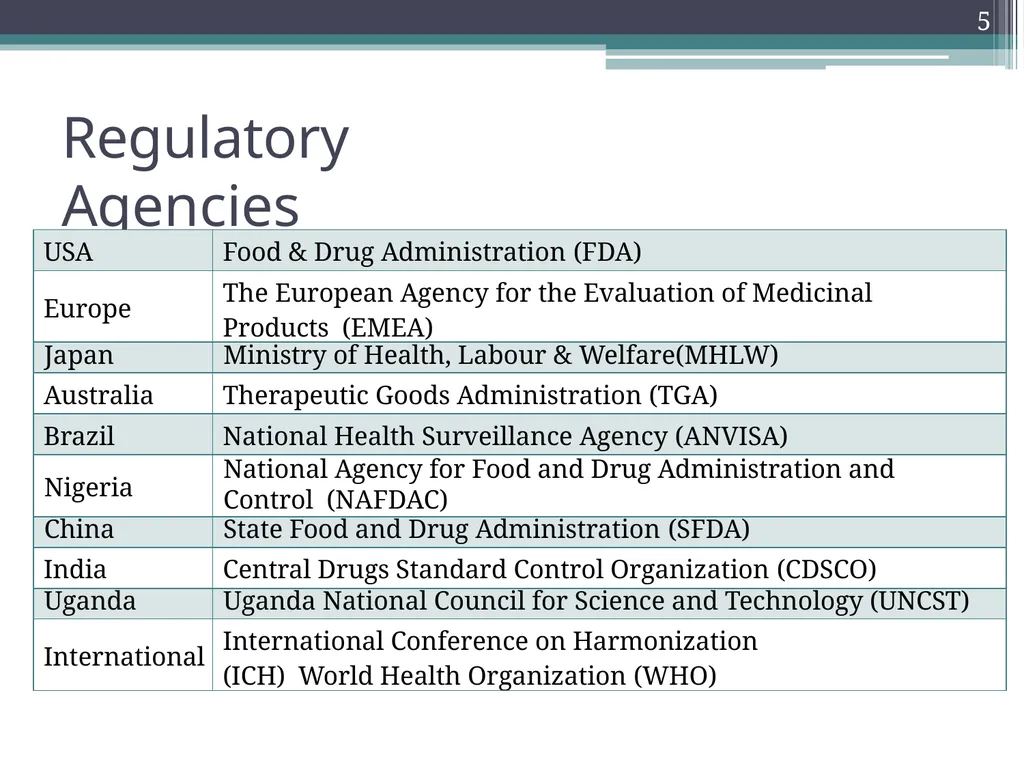
slide5. Regulatory Agencies 5<br>
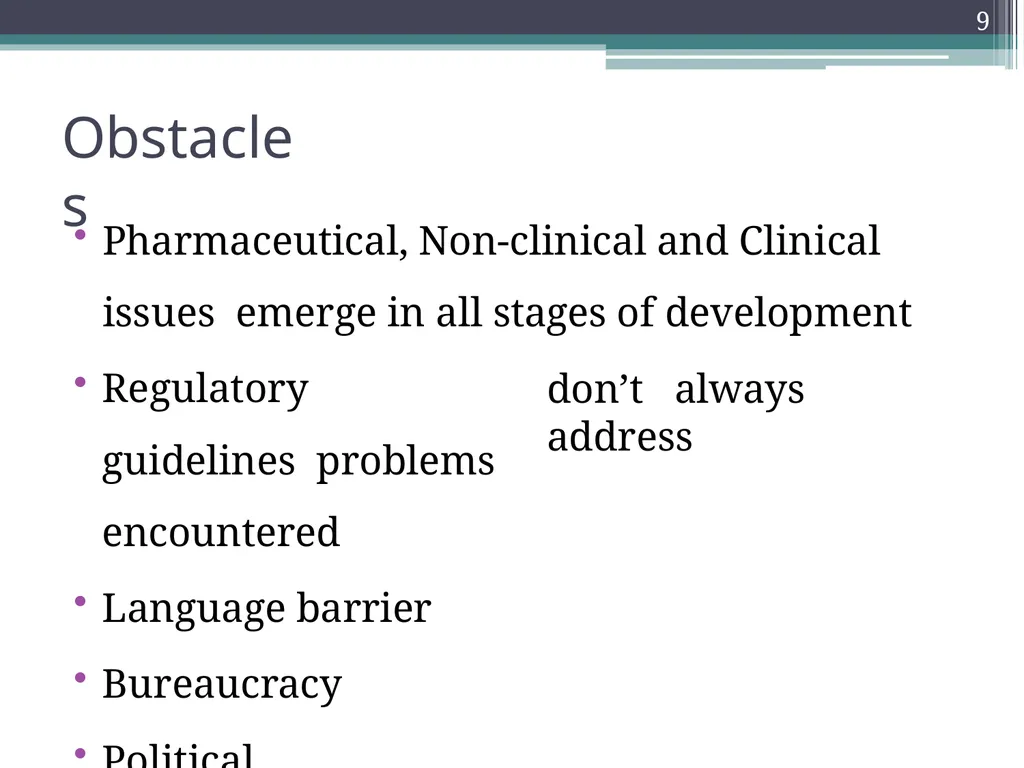
slide6. Obstacles Pharmaceutical, Non-clinical and Clinical issues emerge in all stages of development Regulatory guidelines problems encountered
Language barrier
Bureaucracy
Political environment don’t always address 9<br>
slide7. Registration Procedure of Drugs for International Marketing 10<br>
slide8. Drug Approval Process A regulatory process by which a person/organization/sponsor/innovator gets authorization to launch a drug in the market 11<br>
slide9. Various Stages Application to conduct clinical trials
Conducting clinical trials
Application to marketing authorization of drug
Post-marketing studies 12<br>
slide10. 13<br>
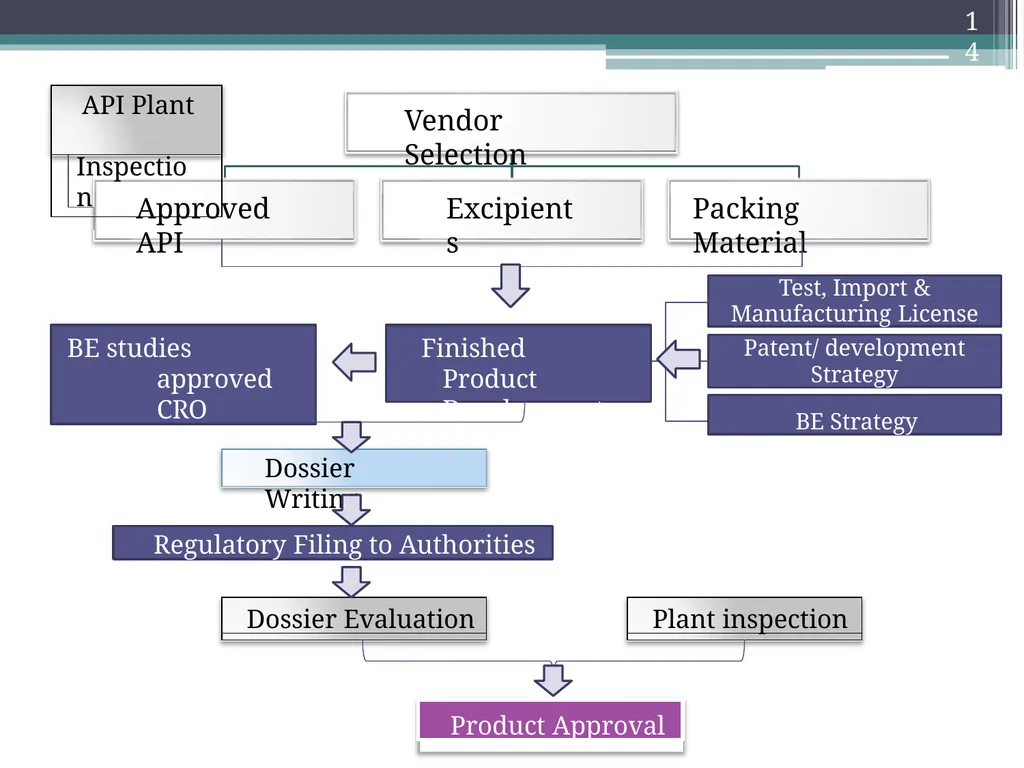
slide11. 14 Vendor Selection Approved API Excipients Packing Material Plant inspection API Plant Inspection BE studies approved CRO Test, Import &
Manufacturing License Patent/ development
Strategy BE Strategy Finished Product Development Dossier Writing Regulatory Filing to Authorities Dossier Evaluation Product Approval<br>
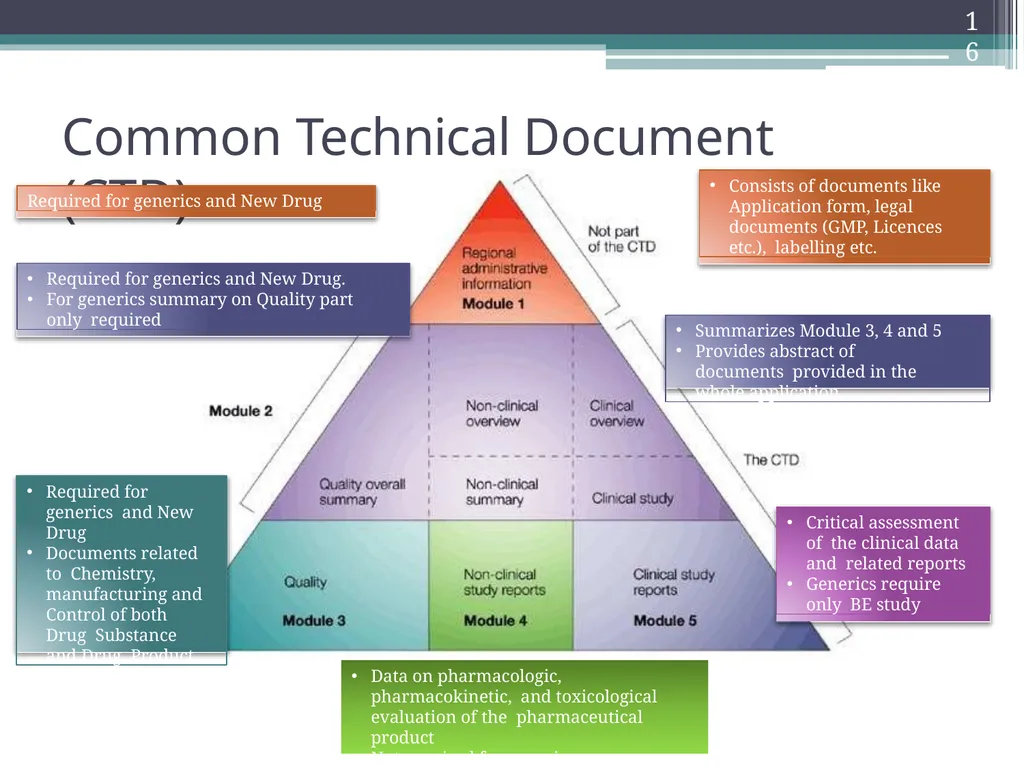
slide12. Common Technical Document (CTD) 16 Required for generics and New Drug Required for generics and New Drug.
For generics summary on Quality part only required Summarizes Module 3, 4 and 5
Provides abstract of documents provided in the whole application Required for generics and New Drug
Documents related to Chemistry, manufacturing and Control of both Drug Substance and Drug Product Data on pharmacologic, pharmacokinetic, and toxicological evaluation of the pharmaceutical product
Not required for generics Critical assessment of the clinical data and related reports
Generics require only BE study Consists of documents like Application form, legal documents (GMP, Licences etc.), labelling etc.<br>
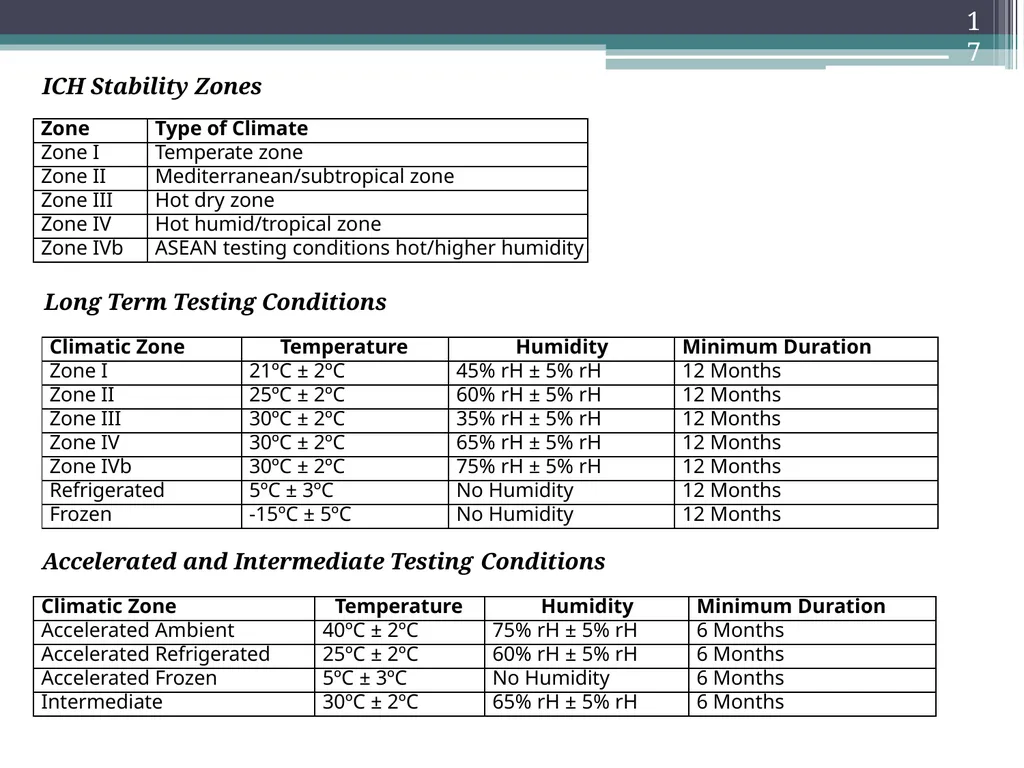
slide13. 17 Accelerated and Intermediate Testing Conditions ICH Stability Zones Long Term Testing Conditions<br>
slide14. COUNTRY SPECIFIC GUIDELINES 18<br>
slide15. Regulatory Agency Food & Drug Administration Main consumer watchdog is FDA's Center for Drug Evaluation in this system
and Research (CDER) 19<br>
slide16. Laws, Regulations, Policies & Procedures The Federal Food, Drug, and Cosmetic Act (1938)
▫ With numerous amendments it is the most extensive law of its kind in the world
Code Of Federal Regulations (CFR)
▫ Section 21 of the CFR contains most regulations pertaining to food and drugs
Manual of Policies and Procedures (MaPPs)
▫ Approved instructions for internal practices and procedures followed by CDER staff to help standardize the new drug review process and other activities 20<br>
slide17. Types of Applications under Sec 505 New Drug Application (NDA)
Abbreviated New Drug Application (ANDA) 21<br>
slide18. New Drug Application (NDA) 22 505 (b) (1) MA for New Chemical Entities (NCE) 505 (b) (2)
MA for changes to an approved drug (Change in dosage form, strength, indication etc.)<br>
slide19. NDA Since 1938, every new drug has been the subject of an
approved NDA before commercialization.
The NDA application is the vehicle through which drug sponsors formally propose that the FDA approve a new pharmaceutical for sale and marketing in the U.S.
The data gathered during the animal studies and human clinical trials of an Investigational New Drug (IND) become part of the NDA.
Generally approval of an NDA is granted within two years (on an average), however, this process can be completed from two months to several years. 23<br>
slide20. Applicable Regulations, Policies & Procedures (NDA) 21CFR Part 314 - Applications for FDA Approval to Market a New Drug or an Antibiotic Drug.
MaPPs
▫ 5015-6:Review of the Same Supplemental Change to More than One NDA or ANDA in More Than One Review Division
▫ 6010.5:NDAs: Filing Review Issues
▫ 6020.8: Action Packages for NDAs and Efficacy Supplements
▫ 6050.1: Refusal to Accept Application for Filing From Applicants in Arrears
▫ 7211.1: Drug Application Approval 501(b) Policy
▫ 7600.6: Requesting and Accepting Non-Archivable Electronic Records for New Drug Applications. 24<br>
slide21. Abbreviated New Drug Application (ANDA) 25 Paragraph I
For the products for which no patent information is available in the orange book Paragraph II Used for the products for which all the applicable patents are expired Paragraph III
Used for the patent products where the applicant confirms that the product will not be placed in the market till such patents are expired Paragraph IV Used for patented products where applicant files the product which does not infringe the patent.
Successful applicant enjoys
6month exclusivity 505 (j)
MA of generics<br>
slide22. ANDA Applications are termed "abbreviated" as preclinical and
clinical data is not required
Reviewers focus on bioequivalence data, chemistry and microbiology data, requests for plant inspection, and drug labelling information.
A generic drug must be comparable to an innovator drug in dosage form, strength, route of administration, quality, performance characteristics and intended use
Once approved, an applicant may manufacture and market the generic drug product to provide a safe, effective, low cost alternative to the American public 26<br>
slide23. Applicable Regulations, Policies & Procedures (ANDA) 21CFR
▫ Part 314 : Applications for FDA Approval to Market a New Drug or an Antibiotic Drug ▫ Part 320 : Bioavailability and Requirements
▫ Part 310 : New Drugs
MaPPs:
▫ Chapter 5200 - Generic Drugs Bioequivalence 27<br>
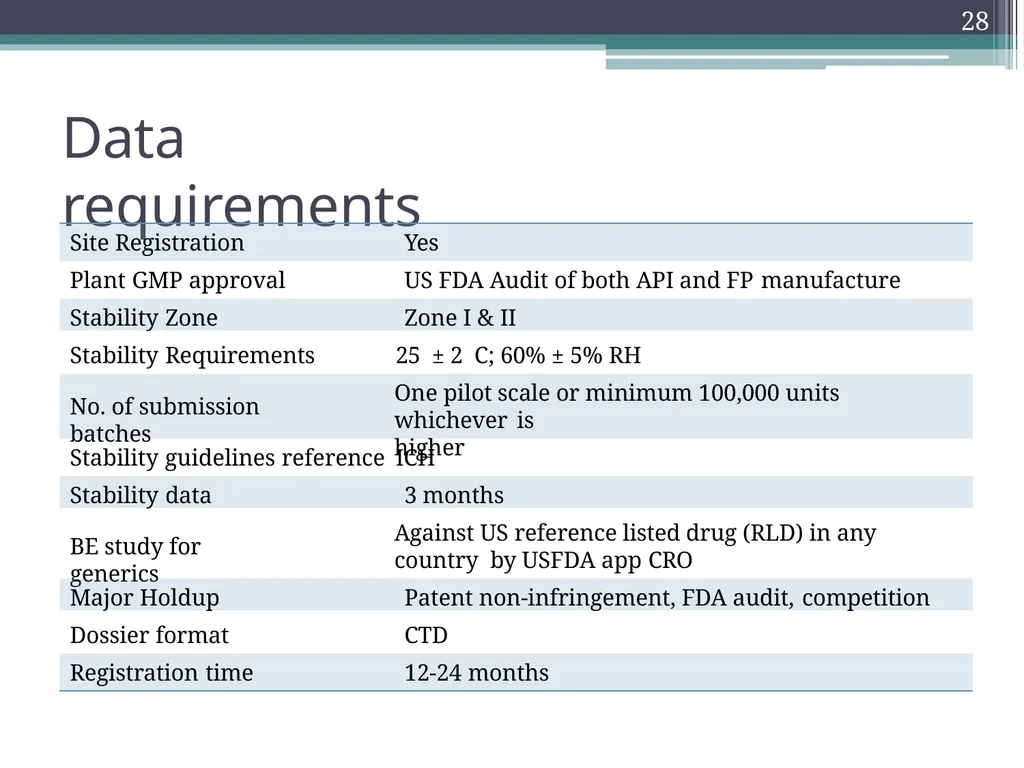
slide24. Data requirements 28 Site Registration Yes
Plant GMP approval US FDA Audit of both API and FP manufacture Stability Zone Zone I & II Stability Requirements 25 ± 2 C; 60% ± 5% RH No. of submission batches One pilot scale or minimum 100,000 units whichever is
higher Stability guidelines reference ICH Stability data 3 months BE study for generics Against US reference listed drug (RLD) in any country by USFDA app CRO Major Holdup Patent non-infringement, FDA audit, competition Dossier format CTD
Registration time 12-24 months<br>
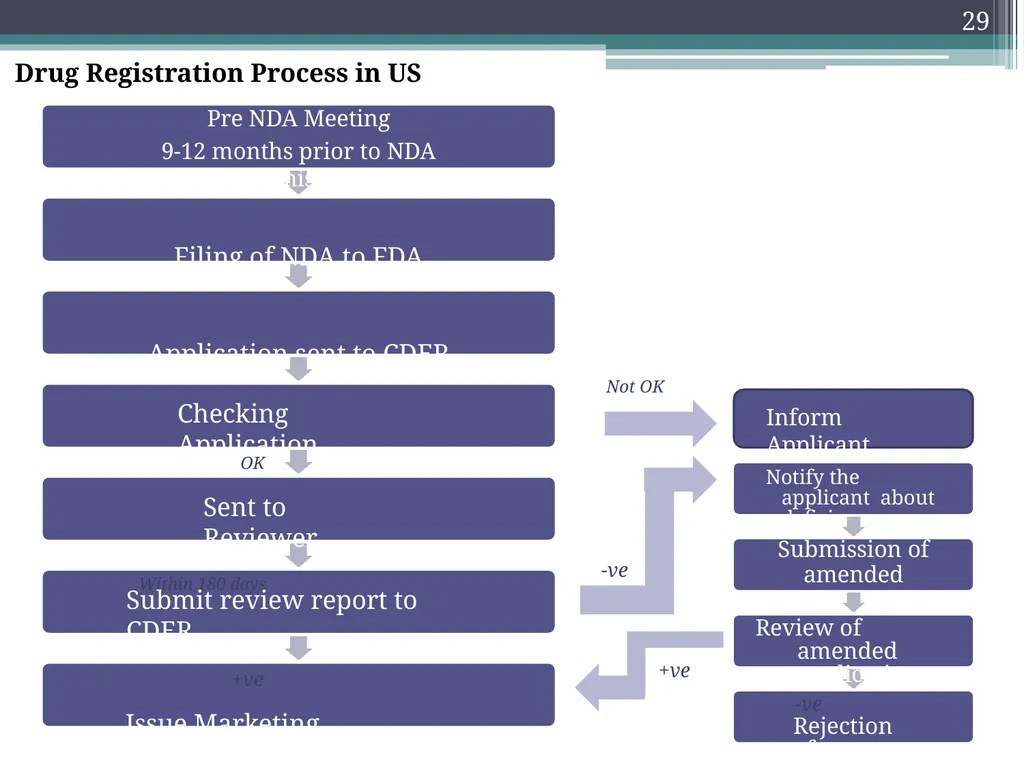
slide25. 29 Drug Registration Process in US Pre NDA Meeting
9-12 months prior to NDA submission
Filing of NDA to FDA
Application sent to CDER Checking Application Sent to Reviewer
Within 180 days Submit review report to CDER
+ve
Issue Marketing Authorization Notify the applicant about deficiency Submission of
amended application Review of amended application
-ve Rejection of Application Inform Applicant Not OK OK -ve +ve<br>
slide26. COUNTRY SPECIFIC GUIDELINES 30<br>
slide27. Regulatory Agencies EMEA Committee for Human Medicinal Products (CHMP) 31<br>
slide28. Types of Procedures Centralized Procedure
Decentralised Procedure
Mutual Recognition Procedure
National Procedure 32<br>
slide29. Centralized Procedure When used
▫ Used for biologic products or other products using high- technology procedures
▫ Products for HIV/AIDS, cancer, diabetes, neurodegenerative disease, auto-immune or other dysfunctions, and viral diseases
▫ Products for orphan conditions
▫ Other new active substances at the request of the applicant
Pros
▫ One application applies to all countries in the EU.
▫ Relatively quick procedure.
▫ A positive outcome is very beneficial to the sponsor.
Cons
▫ A negative outcome will affect access to the entire EU 33<br>
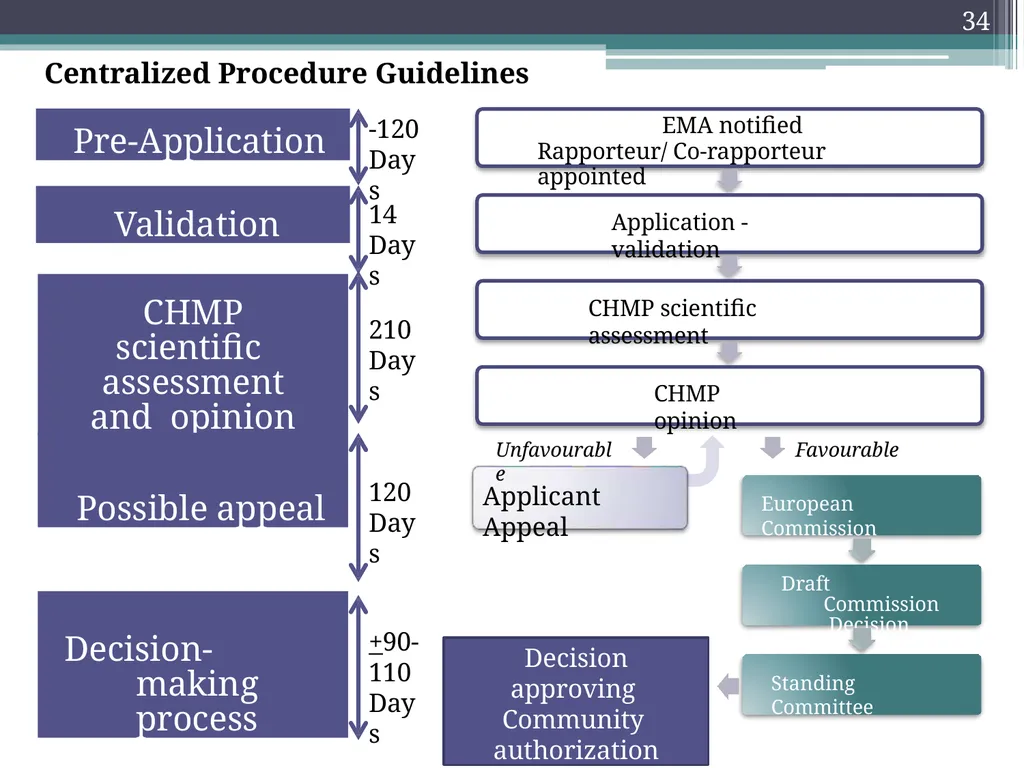
slide30. 34 Pre-Application Validation CHMP scientific assessment and opinion Possible appeal Decision-making process EMA notified Rapporteur/ Co-rapporteur appointed Application - validation CHMP scientific assessment CHMP opinion Applicant Appeal European Commission Draft Commission Decision Standing Committee -120
Days 14
Days 210
Days 120
Days +90-
110
Days Unfavourable Favourable Decision approving Community authorization Centralized Procedure Guidelines<br>
slide31. Decentralised Procedure When used
▫ Used for products that fall outside the scope of the EMA centralized procedure.
Pros
▫ Simultaneous authorization in numerous countries in the EU.
▫ May be more efficient than national authorization since a positive outcome results in numerous country approvals.
▫ Sponsor can select which countries to apply to; does not have to be all EU countries.
Cons
▫ A negative decision on an application may affect numerous countries. 35<br>
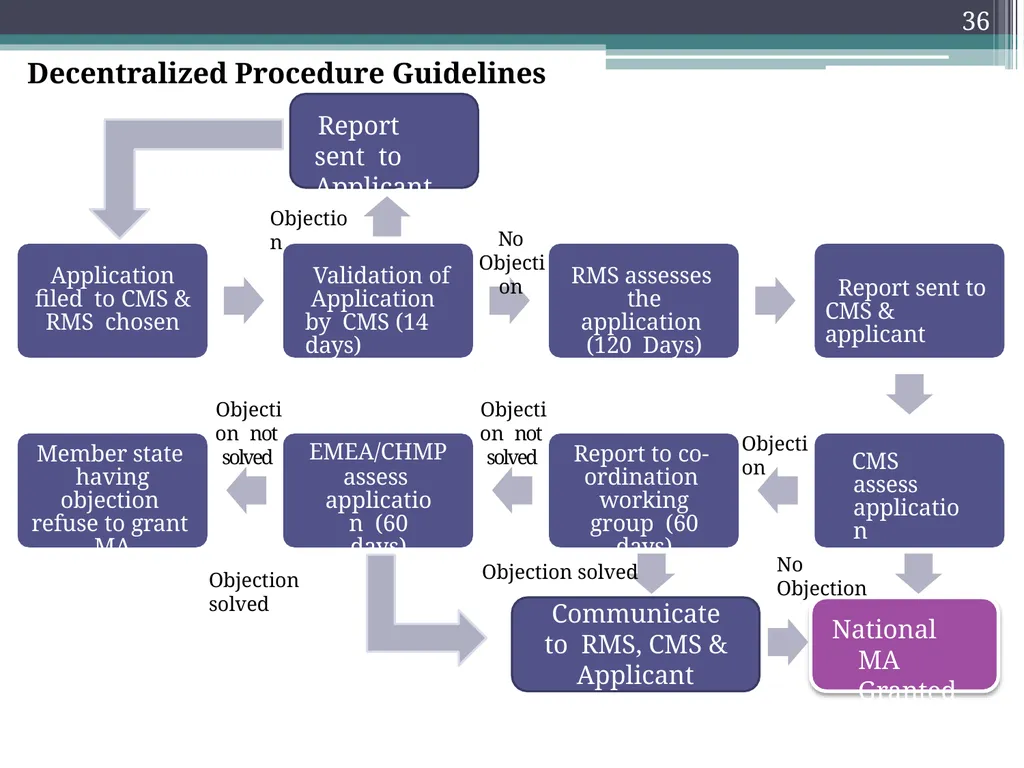
slide32. 36 Application filed to CMS & RMS chosen Validation of Application by CMS (14 days) RMS assesses the application (120 Days) Report sent to CMS & applicant CMS assess application
(90 Days) Report to co- ordination working group (60 days) EMEA/CHMP
assess application (60 days) Member state having objection refuse to grant MA National MA Granted Objection No Objection Objection No
Objection Objection not solved Objection solved
Communicate to RMS, CMS &
Applicant Objection not solved Objection solved Report sent to Applicant Decentralized Procedure Guidelines<br>
slide33. Mutual Recognition Procedure When used
▫ Individual application to one country within the EU for products that fall outside the scope of the EMA centralized procedure.
Pros
▫ Review by one country and other countries accept the decision.
▫ Only one application needs to be submitted.
Cons
▫ Individual national approvals can add significant time
to the process.
▫ A negative outcome can affect numerous countries 37<br>
slide34. Procedure Guidelines Similar to the de-centralized procedure with some differences.
MRP is applicable to medicinal products which have received a marketing authorization in any member state whereas the decentralized procedure is applicable to those products which were never approved in any member states of the European Union.
The evaluation of application by RMS can be taken within 90 days instead of 120 days (in decentralized procedure).
After the grant of marketing authorization, the product can be marketed, which may be called as Phase IV trials, wherein new uses or new populations, long-term effects etc. can be explored. 38<br>
slide35. National Procedure When used
▫ Individual applications to each country within the EU.
▫ Used for products that fall outside the scope of the EMA centralized procedure.
Pros
▫ If application rejected in one country, can still access other EU countries.
Cons
▫ Separate applications required for each country.
▫ Unique requirements and formats may be required. 39<br>
slide36. Procedure Guidelines This type of authorization is granted on country- by-country basis by the competent authorities, in each member state. Each country within the EU procedures for authorizing has its own a marketing application for a new drug.
A sponsor can consult the website of the regulatory agency in each country in which it is interested in obtaining marketing approval to obtain details of the approval process. 40<br>
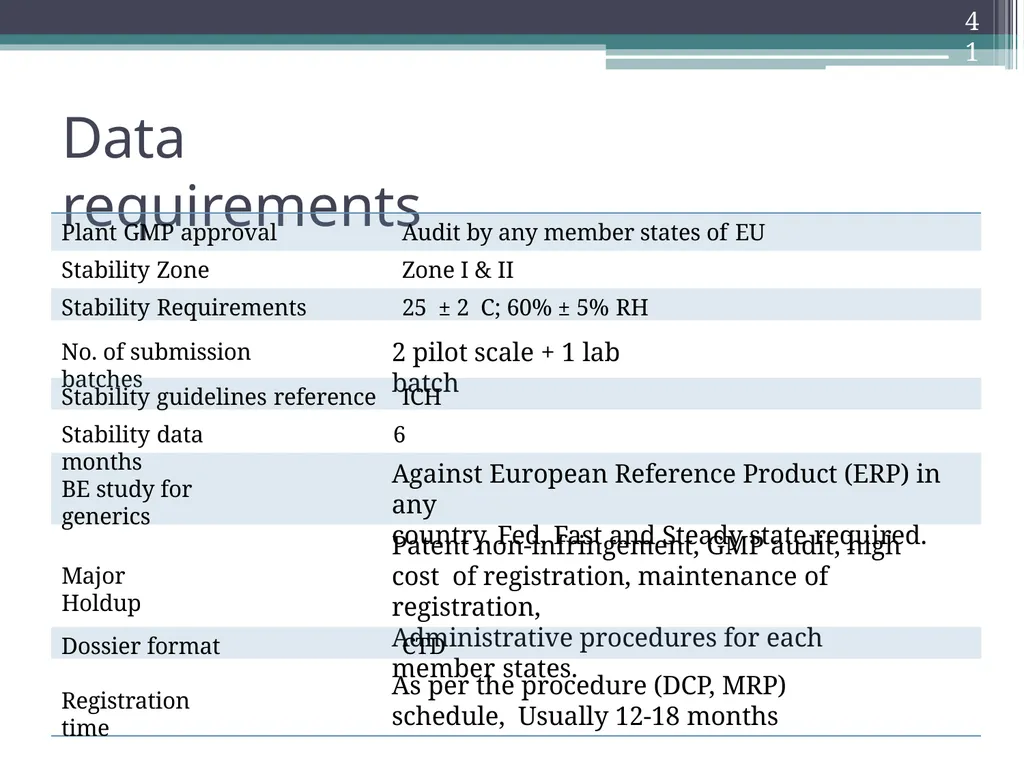
slide37. Data requirements 41 Plant GMP approval Audit by any member states of EU
Stability Zone Zone I & II Stability Requirements 25 ± 2 C; 60% ± 5% RH No. of submission batches 2 pilot scale + 1 lab batch Stability guidelines reference ICH Stability data 6 months BE study for generics Against European Reference Product (ERP) in any
country. Fed, Fast and Steady state required. Major Holdup Patent non-infringement, GMP audit, high cost of registration, maintenance of registration,
Administrative procedures for each member states. Dossier format CTD Registration time As per the procedure (DCP, MRP) schedule, Usually 12-18 months<br>
slide38. COUNTRY SPECIFIC GUIDELINES 42<br>
slide39. Regulatory Agency MHLW Pharmaceutical and Medical Devices (PMDA)
▫ NDA/ANDA Application
▫ Review of GCP/GMP on-site inspection Agency P’ceutical affairs & food (PAFSC)
▫ Consultation on Review Data sanitization council 43<br>
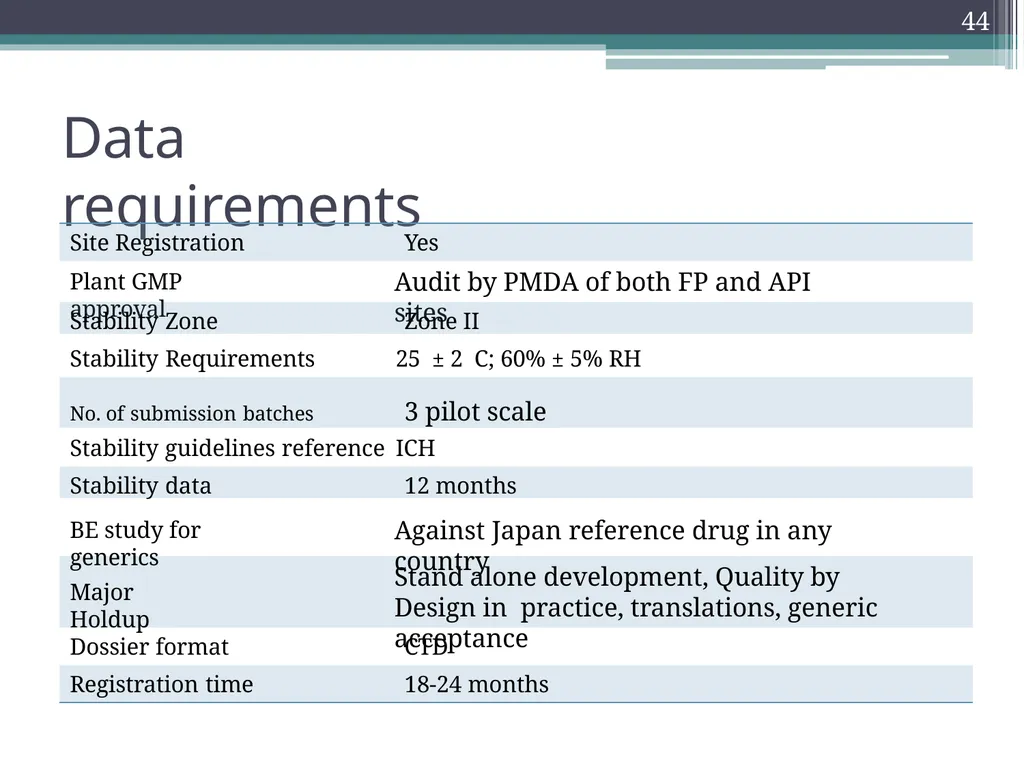
slide40. Data requirements 44 Site Registration Yes Plant GMP approval Audit by PMDA of both FP and API sites Stability Zone Zone II Stability Requirements 25 ± 2 C; 60% ± 5% RH No. of submission batches 3 pilot scale Stability guidelines reference ICH Stability data 12 months BE study for generics Against Japan reference drug in any country Major Holdup Stand alone development, Quality by Design in practice, translations, generic acceptance Dossier format CTD
Registration time 18-24 months<br>
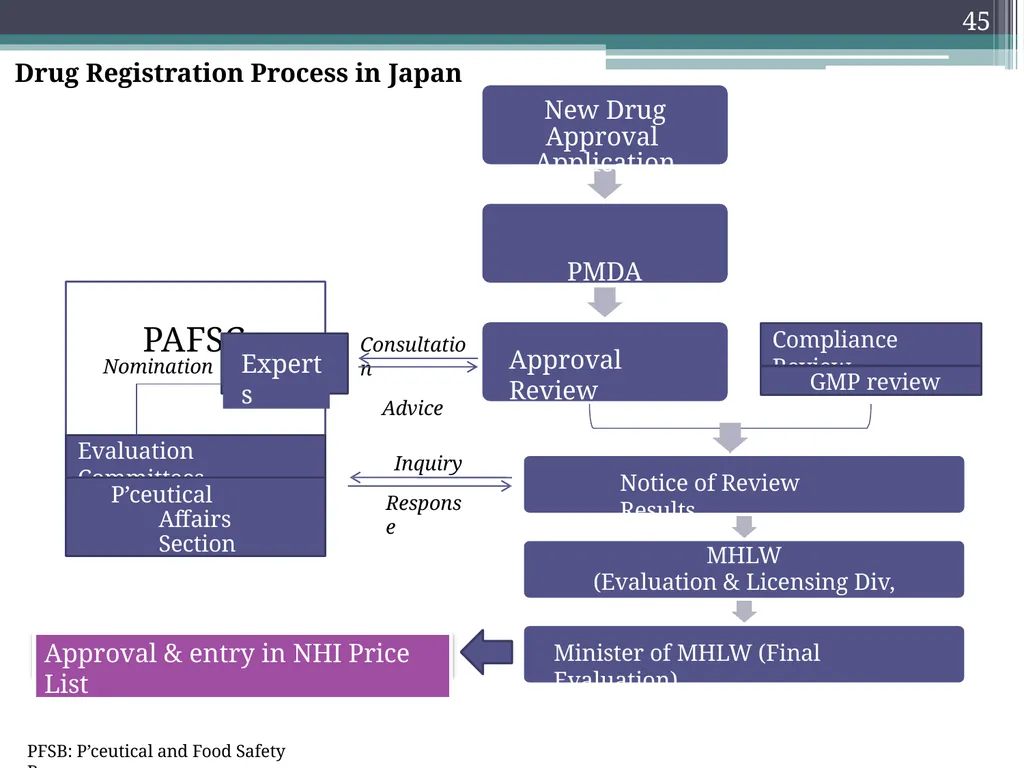
slide41. 45 Approval Review Compliance Review GMP review Drug Registration Process in Japan
New Drug Approval Application
PMDA
PAFSC Evaluation Committees P’ceutical Affairs Section Experts Nomination Consultation
Advice Inquiry
Response Notice of Review Results MHLW
(Evaluation & Licensing Div, PFSB) Minister of MHLW (Final Evaluation) PFSB: P’ceutical and Food Safety Bureau Approval & entry in NHI Price List<br>
slide42. COUNTRY SPECIFIC GUIDELINES 46<br>
slide43. Regulatory Agency The TGA regulates therapeutic goods through:
▫ Pre-market assessment
▫ Post-market monitoring and enforcement of standards
▫ Licensing of Australian manufacturers and verifying overseas manufacturers' compliance with the same standards as their Australian counterparts
Australian Drug Evaluation Committee (ADEC)
▫ Gives advice for new medicines
Drug Safety and Evaluation Branch (DSEB)
▫ NDA/ANDA Application 47<br>
slide44. Guidelines An application is submitted to TGA to register the drug in Australian Register of Therapeutic Goods (ARTG) after the completion of clinical trials.
The application consists of data to support the quality, safety and efficacy of the product for its intended use.
The application is assessed (on an administrative level) to make sure for compliance with basic guidelines and further evaluated by different sections and advice can also be sought on key issues to take final decision.
A company can make comments on the evaluation report, if necessary. 48<br>
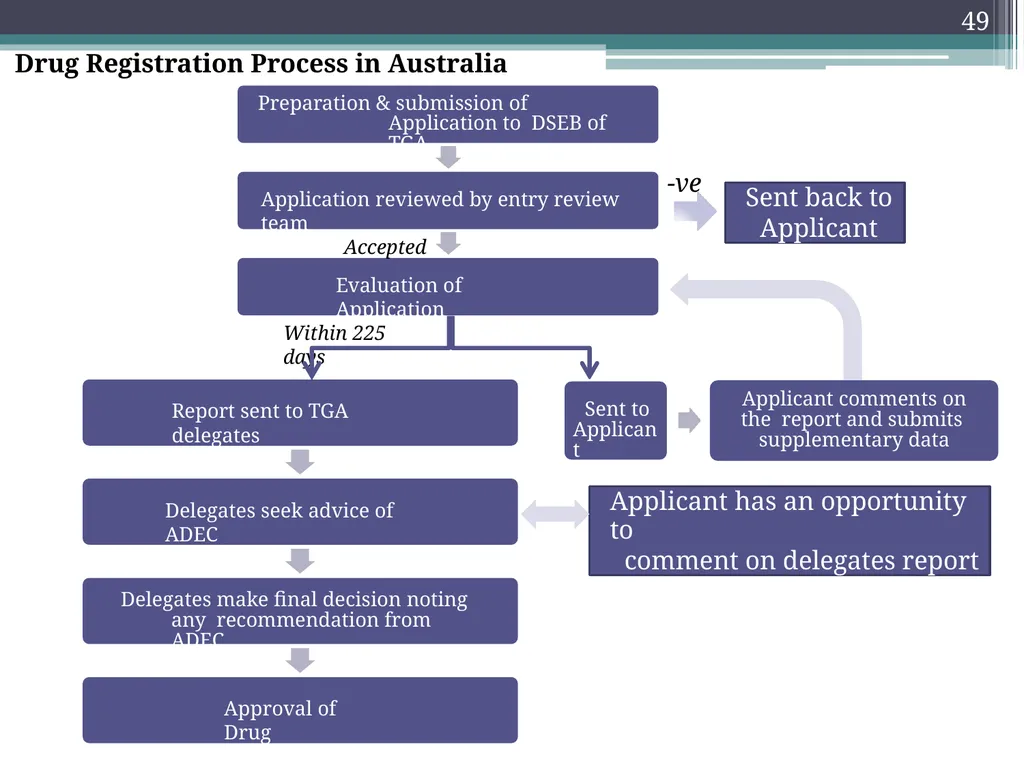
slide45. 49 Preparation & submission of Application to DSEB of TGA Application reviewed by entry review team Accepted
Evaluation of Application -ve Sent back to
Applicant Within 225 days Report sent to TGA delegates Delegates seek advice of ADEC Delegates make final decision noting any recommendation from ADEC Approval of Drug Sent to Applicant Applicant comments on the report and submits supplementary data Applicant has an opportunity to
comment on delegates report Drug Registration Process in Australia<br>
slide46. COUNTRY SPECIFIC GUIDELINES 50<br>
slide47. Regulatory Agency State Food and Drug Administration (SFDA) Provincial Drug Administration Authorities (PDAAs)
▫ Organize the works of the formal review of submitted materials
Centre for Drug Evaluation (CDE)
▫ Evaluate the submitted information 51<br>
slide48. Drug Approval Process After formal review, the PDAAs send the qualified applications to the SFDA for further review.
The import drug registration application should be directly submitted to SFDA by the applicant.
SFDA’s Department of Drug Registration carefully reviews the completeness of the submitted materials, files the qualified applications and transmits all the materials of qualified applications to the directly attached to SFDA.
CDE determine whether the safety and effectiveness information submitted for a new drug are adequate for manufacturing and marketing approval and send the report of review to SFDA.
SFDA carefully considers the recommendations of CDE and makes a decision whether or not the drug registration application can be approved and issues the certificate of drug approval and drug approval number to the qualified applicant. 52<br>
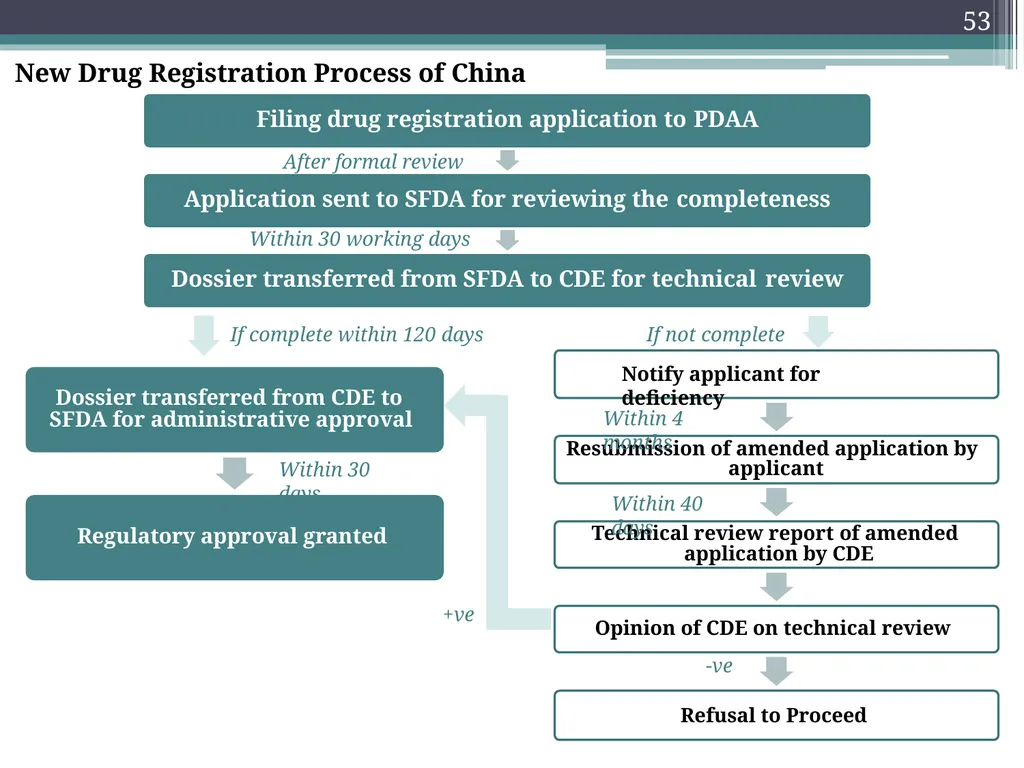
slide49. 53 Dossier transferred from CDE to SFDA for administrative approval Regulatory approval granted Within 30 days Resubmission of amended application by applicant Technical review report of amended application by CDE Opinion of CDE on technical review Filing drug registration application to PDAA
After formal review
Application sent to SFDA for reviewing the completeness
Within 30 working days
Dossier transferred from SFDA to CDE for technical review
If complete within 120 days If not complete
Notify applicant for deficiency Within 4 months Within 40 days +ve -ve
Refusal to Proceed New Drug Registration Process of China<br>
slide50. COUNTRY SPECIFIC GUIDELINES 54<br>
slide51. Markets Brazil, Mexico, Venezuela , Ecuador and Colombia are some of the major pharmaceutical markets
Brazil and Mexico considered Regulated Markets
ANVISA : Stringent norms comparable to USFDA
Brazil : As per ANVISAs norms, CROs used for BE studies have to be approved and certified by the Brazilian authority.
For BE studies, innovator product manufactured in Brazil has to be used for comparison
ANVISA has implemented generic law to facilitate generic
market entry into the country
Mexico: Extensive analytical raw data of 3 stability batches.
Venezuela : Copies of source data 55<br>
slide52. Drug Regulatory Requirements - Technical Manufacturing Formula and Justification
Manufacturing Process
Process Validation
Batch Numbering
Batch Records
Stability Studies (Source Data)
▫ Accelerated
▫ Long Term 56<br>
slide53. Drug Regulatory Requirements - Legal Free Sale Certificate / Product Permission
GMP Certificate
Manufacturing License
Power of Attorney / Contract
Trademark Registration
Contract with local quality control facility 57<br>
slide54. Critical Success Factors US FDA / UK MCA / TGA approved facilities
Strong Regulatory Support
Good Distribution Set up
Price Competitiveness
Good Marketing Support
Strong group for registering products in place
Expansion in new markets and new products under feasibility studies 58<br>
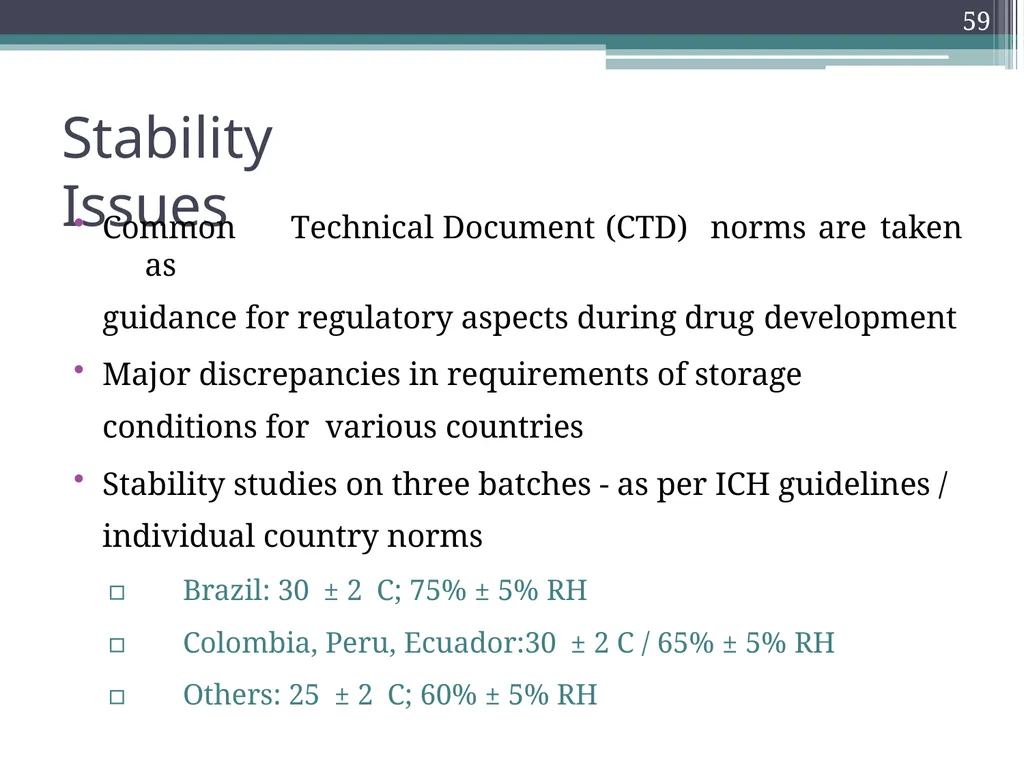
slide55. Stability Issues Common Technical Document (CTD) norms are taken as
guidance for regulatory aspects during drug development
Major discrepancies in requirements of storage conditions for various countries
Stability studies on three batches - as per ICH guidelines / individual country norms
▫ Brazil: 30 ± 2 C; 75% ± 5% RH
▫ Colombia, Peru, Ecuador:30 ± 2C / 65% ± 5% RH
▫ Others: 25 ± 2 C; 60% ± 5% RH 59<br>
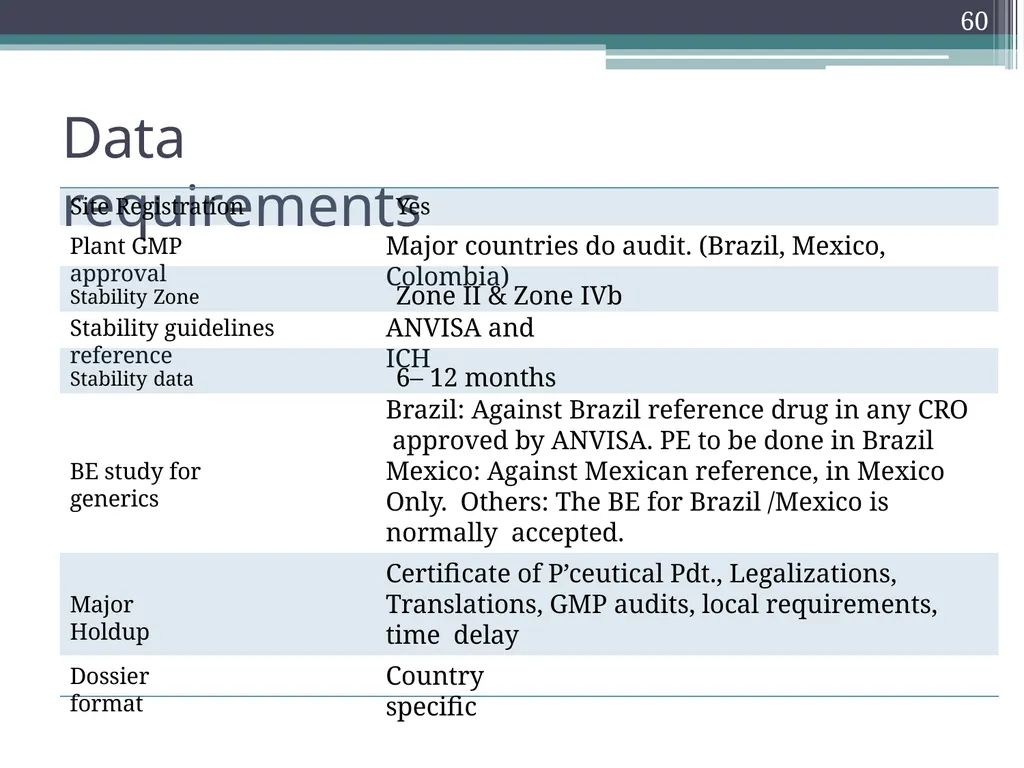
slide56. Data requirements 60 Site Registration Yes Plant GMP approval Major countries do audit. (Brazil, Mexico, Colombia) Stability Zone Zone II & Zone IVb Stability guidelines reference ANVISA and ICH Stability data 6– 12 months BE study for generics Brazil: Against Brazil reference drug in any CRO approved by ANVISA. PE to be done in Brazil Mexico: Against Mexican reference, in Mexico Only. Others: The BE for Brazil /Mexico is normally accepted. Major Holdup Certificate of P’ceutical Pdt., Legalizations, Translations, GMP audits, local requirements, time delay Dossier format Country specific<br>
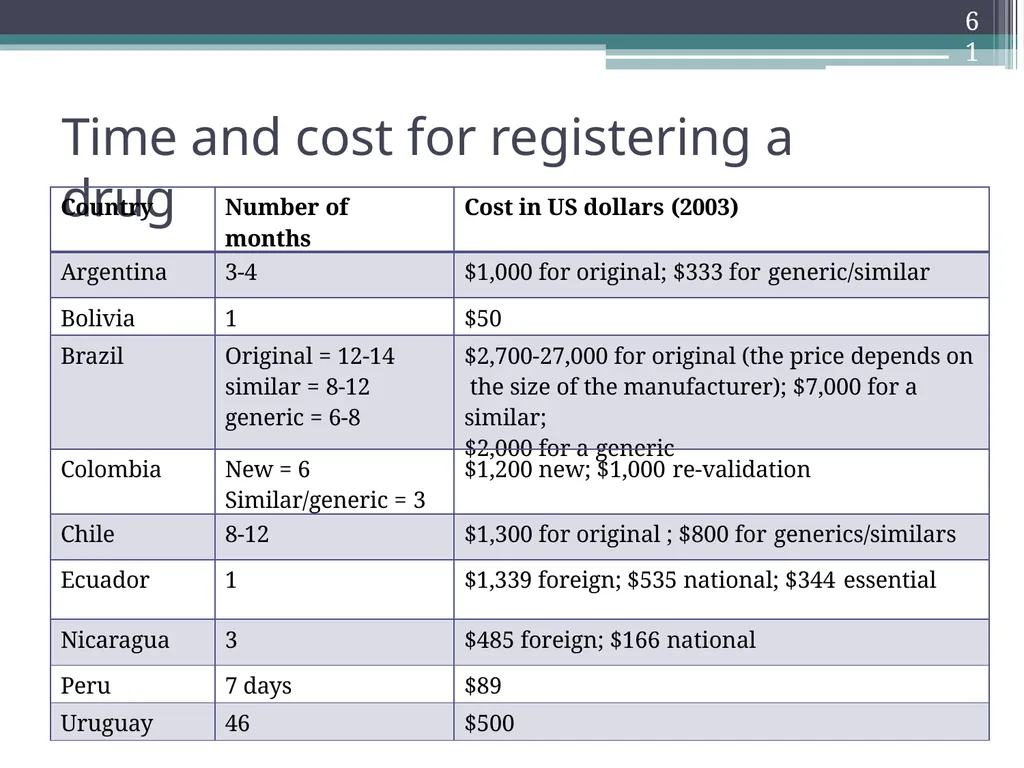
slide57. Time and cost for registering a drug 61<br>
slide58. COUNTRY SPECIFIC GUIDELINES 62<br>
slide59. Guidelines Russia has its own, unique regulatory system which does not correspond to U.S. or EU practices.
Cultural and language barriers often become a challenge to foreign companies attempting to register pharmaceuticals by themselves
Registration procedure is also quite complicated and the documents tend to be modified due to constant changes in the regulatory requirements.
Before exporting a pharmaceutical product to Russia it has to be registered at the Ministry of Health of the Russian Federation; Department for registration of pharmaceuticals of the Federal Service for Health Control 63<br>
slide60. Registration Procedure The best way to accomplish registration of a pharmaceutical product in Russia is trough one of the following types of companies:
▫ Company incorporated in Russia and belonging to the Indian parent company
▫ Russian distributor/authorized agent
▫ Consulting company 64<br>
slide61. Documentation For the registration procedure a company has to submit the following:
▫ General documents
▫ Certificates
▫ Information and test reports
▫ Samples and package 65<br>
slide62. General documents Application for the State registration of a pharmaceutical which includes the name of the pharmaceutical preparation, the name and the contact information of the manufacturer
Name of the pharmaceutical preparation, including international non-proprietary name, scientific name in Latin, trade name and main synonyms
List of active ingredients and components
Recommended dosage, instruction for use
Description of the drug and its packaging, shelf life and
storage conditions
Power of Attorney issued by the manufacturer to the authorized company for carrying out registration procedure (notarized original with apostil) 66<br>
slide63. Certificates Copy of Free Sales Certificate (must be notarized and apostilled)
Copy of the license of pharmaceutical manufacture (must be notarized and apostilled)
Copy of GMP certificate (must be notarized and apostilled)
Copy of Certificate of manufacturer registration in their own country (must be notarized and apostilled)
Original of Certificate of analysis of the drug and its active substance (must
be signed and stamped by manufacturer)
Copy of Certificate of trade mark (must be signed and stamped by the manufacturer)
Information of registration of the drug in the country of the manufacture
and in other countries
A certificate of analysis of the drug in the country of origin of the manufacturer, GMP certificate and information on registration of the drug in the foreign country. 67<br>
slide64. Information and test reports Summary of the method of the drug manufacturing (must be signed and stamped by manufacturer)
Complete description of the quantitative and qualitative control methods with references to the pharmacopoeia and specification (must be signed and stamped by manufacturer)
Stability data of three drug series – by date
Patterns of the spectrums and chromatograms of the drug
Report of the pharmacological (specific) activity study substantiated the indications for use which are formed and described in the instruction
Test report of the drug toxicity (acute, sub acute, sub chronic, chronic toxicity) 68<br>
slide65. Information and test reports Test report of the specific influences (cancerogenity, mutagenic and teratogenic effects, embryotoxicity, allergic and local-irritative effects)
Clinical trial report of the medicine usage in clinic (the information which concerns only the drug that is produced by this manufacturer)
Copies of publications of the medicine usage in clinics after its registration in the country of origin (the information which concerns only the drug that is produced by this manufacturer)
Report of pharmacokinetics of the pharmaceutical study and its
bioequivalence to the original drug
Summary information of the side effects, in comparison with other analogous medicines, used at the same indications
Instruction for use (must be signed and stamped by the manufacturer) 69<br>
slide66. Samples and package Information of the material used for package: Certificates of the packaging materials (must be signed and stamped by the manufacturer)
Colour design of internal and external packages (Original and Russian version)
Standard samples of the active substance for quality control
Standard and referenced samples of the drug for the binding examination of quality (should be in the standard package) 70<br>
slide67. Approval Procedure The Ministry of Health determines whether these approvals are sufficient for an exemption of the drug from clinical and other testing in Russia before issuing a registration certificate.
Officially Russia does not recognize FDA and EU certificates.
According to the Law on Medicines, a fee in form of a state duty is to be paid for state registration and the registration should take no longer than six month. 71<br>
Drug Regulatory Affairs 1<br>
slide2. Regulatory structure The pharmaceutical industry is one of the highly regulated industries with various structures of drug regulation
▫ Drug laws
▫ Drug regulatory agencies
▫ Drug evaluation boards
▫ Quality control laboratories
▫ Drug information centers 2<br>
slide3. What is the need? Drug laws provide the basis for drug regulation
Regulatory tools such as standards and guidelines equip authorities with the practical means of implementing those laws
Regulatory Agencies monitor the implementation of these laws 3<br>
slide4. Regulatory Scenario Regulated Markets Emerging Markets US
Canada
EU-27 states
Japan
Australia
New Zealand Latin America
Eastern Europe
CIS
Africa
Asia Pacific
Gulf countries 4<br>
slide5. Regulatory Agencies 5<br>
slide6. Obstacles Pharmaceutical, Non-clinical and Clinical issues emerge in all stages of development Regulatory guidelines problems encountered
Language barrier
Bureaucracy
Political environment don’t always address 9<br>
slide7. Registration Procedure of Drugs for International Marketing 10<br>
slide8. Drug Approval Process A regulatory process by which a person/organization/sponsor/innovator gets authorization to launch a drug in the market 11<br>
slide9. Various Stages Application to conduct clinical trials
Conducting clinical trials
Application to marketing authorization of drug
Post-marketing studies 12<br>
slide10. 13<br>
slide11. 14 Vendor Selection Approved API Excipients Packing Material Plant inspection API Plant Inspection BE studies approved CRO Test, Import &
Manufacturing License Patent/ development
Strategy BE Strategy Finished Product Development Dossier Writing Regulatory Filing to Authorities Dossier Evaluation Product Approval<br>
slide12. Common Technical Document (CTD) 16 Required for generics and New Drug Required for generics and New Drug.
For generics summary on Quality part only required Summarizes Module 3, 4 and 5
Provides abstract of documents provided in the whole application Required for generics and New Drug
Documents related to Chemistry, manufacturing and Control of both Drug Substance and Drug Product Data on pharmacologic, pharmacokinetic, and toxicological evaluation of the pharmaceutical product
Not required for generics Critical assessment of the clinical data and related reports
Generics require only BE study Consists of documents like Application form, legal documents (GMP, Licences etc.), labelling etc.<br>
slide13. 17 Accelerated and Intermediate Testing Conditions ICH Stability Zones Long Term Testing Conditions<br>
slide14. COUNTRY SPECIFIC GUIDELINES 18<br>
slide15. Regulatory Agency Food & Drug Administration Main consumer watchdog is FDA's Center for Drug Evaluation in this system
and Research (CDER) 19<br>
slide16. Laws, Regulations, Policies & Procedures The Federal Food, Drug, and Cosmetic Act (1938)
▫ With numerous amendments it is the most extensive law of its kind in the world
Code Of Federal Regulations (CFR)
▫ Section 21 of the CFR contains most regulations pertaining to food and drugs
Manual of Policies and Procedures (MaPPs)
▫ Approved instructions for internal practices and procedures followed by CDER staff to help standardize the new drug review process and other activities 20<br>
slide17. Types of Applications under Sec 505 New Drug Application (NDA)
Abbreviated New Drug Application (ANDA) 21<br>
slide18. New Drug Application (NDA) 22 505 (b) (1) MA for New Chemical Entities (NCE) 505 (b) (2)
MA for changes to an approved drug (Change in dosage form, strength, indication etc.)<br>
slide19. NDA Since 1938, every new drug has been the subject of an
approved NDA before commercialization.
The NDA application is the vehicle through which drug sponsors formally propose that the FDA approve a new pharmaceutical for sale and marketing in the U.S.
The data gathered during the animal studies and human clinical trials of an Investigational New Drug (IND) become part of the NDA.
Generally approval of an NDA is granted within two years (on an average), however, this process can be completed from two months to several years. 23<br>
slide20. Applicable Regulations, Policies & Procedures (NDA) 21CFR Part 314 - Applications for FDA Approval to Market a New Drug or an Antibiotic Drug.
MaPPs
▫ 5015-6:Review of the Same Supplemental Change to More than One NDA or ANDA in More Than One Review Division
▫ 6010.5:NDAs: Filing Review Issues
▫ 6020.8: Action Packages for NDAs and Efficacy Supplements
▫ 6050.1: Refusal to Accept Application for Filing From Applicants in Arrears
▫ 7211.1: Drug Application Approval 501(b) Policy
▫ 7600.6: Requesting and Accepting Non-Archivable Electronic Records for New Drug Applications. 24<br>
slide21. Abbreviated New Drug Application (ANDA) 25 Paragraph I
For the products for which no patent information is available in the orange book Paragraph II Used for the products for which all the applicable patents are expired Paragraph III
Used for the patent products where the applicant confirms that the product will not be placed in the market till such patents are expired Paragraph IV Used for patented products where applicant files the product which does not infringe the patent.
Successful applicant enjoys
6month exclusivity 505 (j)
MA of generics<br>
slide22. ANDA Applications are termed "abbreviated" as preclinical and
clinical data is not required
Reviewers focus on bioequivalence data, chemistry and microbiology data, requests for plant inspection, and drug labelling information.
A generic drug must be comparable to an innovator drug in dosage form, strength, route of administration, quality, performance characteristics and intended use
Once approved, an applicant may manufacture and market the generic drug product to provide a safe, effective, low cost alternative to the American public 26<br>
slide23. Applicable Regulations, Policies & Procedures (ANDA) 21CFR
▫ Part 314 : Applications for FDA Approval to Market a New Drug or an Antibiotic Drug ▫ Part 320 : Bioavailability and Requirements
▫ Part 310 : New Drugs
MaPPs:
▫ Chapter 5200 - Generic Drugs Bioequivalence 27<br>
slide24. Data requirements 28 Site Registration Yes
Plant GMP approval US FDA Audit of both API and FP manufacture Stability Zone Zone I & II Stability Requirements 25 ± 2 C; 60% ± 5% RH No. of submission batches One pilot scale or minimum 100,000 units whichever is
higher Stability guidelines reference ICH Stability data 3 months BE study for generics Against US reference listed drug (RLD) in any country by USFDA app CRO Major Holdup Patent non-infringement, FDA audit, competition Dossier format CTD
Registration time 12-24 months<br>
slide25. 29 Drug Registration Process in US Pre NDA Meeting
9-12 months prior to NDA submission
Filing of NDA to FDA
Application sent to CDER Checking Application Sent to Reviewer
Within 180 days Submit review report to CDER
+ve
Issue Marketing Authorization Notify the applicant about deficiency Submission of
amended application Review of amended application
-ve Rejection of Application Inform Applicant Not OK OK -ve +ve<br>
slide26. COUNTRY SPECIFIC GUIDELINES 30<br>
slide27. Regulatory Agencies EMEA Committee for Human Medicinal Products (CHMP) 31<br>
slide28. Types of Procedures Centralized Procedure
Decentralised Procedure
Mutual Recognition Procedure
National Procedure 32<br>
slide29. Centralized Procedure When used
▫ Used for biologic products or other products using high- technology procedures
▫ Products for HIV/AIDS, cancer, diabetes, neurodegenerative disease, auto-immune or other dysfunctions, and viral diseases
▫ Products for orphan conditions
▫ Other new active substances at the request of the applicant
Pros
▫ One application applies to all countries in the EU.
▫ Relatively quick procedure.
▫ A positive outcome is very beneficial to the sponsor.
Cons
▫ A negative outcome will affect access to the entire EU 33<br>
slide30. 34 Pre-Application Validation CHMP scientific assessment and opinion Possible appeal Decision-making process EMA notified Rapporteur/ Co-rapporteur appointed Application - validation CHMP scientific assessment CHMP opinion Applicant Appeal European Commission Draft Commission Decision Standing Committee -120
Days 14
Days 210
Days 120
Days +90-
110
Days Unfavourable Favourable Decision approving Community authorization Centralized Procedure Guidelines<br>
slide31. Decentralised Procedure When used
▫ Used for products that fall outside the scope of the EMA centralized procedure.
Pros
▫ Simultaneous authorization in numerous countries in the EU.
▫ May be more efficient than national authorization since a positive outcome results in numerous country approvals.
▫ Sponsor can select which countries to apply to; does not have to be all EU countries.
Cons
▫ A negative decision on an application may affect numerous countries. 35<br>
slide32. 36 Application filed to CMS & RMS chosen Validation of Application by CMS (14 days) RMS assesses the application (120 Days) Report sent to CMS & applicant CMS assess application
(90 Days) Report to co- ordination working group (60 days) EMEA/CHMP
assess application (60 days) Member state having objection refuse to grant MA National MA Granted Objection No Objection Objection No
Objection Objection not solved Objection solved
Communicate to RMS, CMS &
Applicant Objection not solved Objection solved Report sent to Applicant Decentralized Procedure Guidelines<br>
slide33. Mutual Recognition Procedure When used
▫ Individual application to one country within the EU for products that fall outside the scope of the EMA centralized procedure.
Pros
▫ Review by one country and other countries accept the decision.
▫ Only one application needs to be submitted.
Cons
▫ Individual national approvals can add significant time
to the process.
▫ A negative outcome can affect numerous countries 37<br>
slide34. Procedure Guidelines Similar to the de-centralized procedure with some differences.
MRP is applicable to medicinal products which have received a marketing authorization in any member state whereas the decentralized procedure is applicable to those products which were never approved in any member states of the European Union.
The evaluation of application by RMS can be taken within 90 days instead of 120 days (in decentralized procedure).
After the grant of marketing authorization, the product can be marketed, which may be called as Phase IV trials, wherein new uses or new populations, long-term effects etc. can be explored. 38<br>
slide35. National Procedure When used
▫ Individual applications to each country within the EU.
▫ Used for products that fall outside the scope of the EMA centralized procedure.
Pros
▫ If application rejected in one country, can still access other EU countries.
Cons
▫ Separate applications required for each country.
▫ Unique requirements and formats may be required. 39<br>
slide36. Procedure Guidelines This type of authorization is granted on country- by-country basis by the competent authorities, in each member state. Each country within the EU procedures for authorizing has its own a marketing application for a new drug.
A sponsor can consult the website of the regulatory agency in each country in which it is interested in obtaining marketing approval to obtain details of the approval process. 40<br>
slide37. Data requirements 41 Plant GMP approval Audit by any member states of EU
Stability Zone Zone I & II Stability Requirements 25 ± 2 C; 60% ± 5% RH No. of submission batches 2 pilot scale + 1 lab batch Stability guidelines reference ICH Stability data 6 months BE study for generics Against European Reference Product (ERP) in any
country. Fed, Fast and Steady state required. Major Holdup Patent non-infringement, GMP audit, high cost of registration, maintenance of registration,
Administrative procedures for each member states. Dossier format CTD Registration time As per the procedure (DCP, MRP) schedule, Usually 12-18 months<br>
slide38. COUNTRY SPECIFIC GUIDELINES 42<br>
slide39. Regulatory Agency MHLW Pharmaceutical and Medical Devices (PMDA)
▫ NDA/ANDA Application
▫ Review of GCP/GMP on-site inspection Agency P’ceutical affairs & food (PAFSC)
▫ Consultation on Review Data sanitization council 43<br>
slide40. Data requirements 44 Site Registration Yes Plant GMP approval Audit by PMDA of both FP and API sites Stability Zone Zone II Stability Requirements 25 ± 2 C; 60% ± 5% RH No. of submission batches 3 pilot scale Stability guidelines reference ICH Stability data 12 months BE study for generics Against Japan reference drug in any country Major Holdup Stand alone development, Quality by Design in practice, translations, generic acceptance Dossier format CTD
Registration time 18-24 months<br>
slide41. 45 Approval Review Compliance Review GMP review Drug Registration Process in Japan
New Drug Approval Application
PMDA
PAFSC Evaluation Committees P’ceutical Affairs Section Experts Nomination Consultation
Advice Inquiry
Response Notice of Review Results MHLW
(Evaluation & Licensing Div, PFSB) Minister of MHLW (Final Evaluation) PFSB: P’ceutical and Food Safety Bureau Approval & entry in NHI Price List<br>
slide42. COUNTRY SPECIFIC GUIDELINES 46<br>
slide43. Regulatory Agency The TGA regulates therapeutic goods through:
▫ Pre-market assessment
▫ Post-market monitoring and enforcement of standards
▫ Licensing of Australian manufacturers and verifying overseas manufacturers' compliance with the same standards as their Australian counterparts
Australian Drug Evaluation Committee (ADEC)
▫ Gives advice for new medicines
Drug Safety and Evaluation Branch (DSEB)
▫ NDA/ANDA Application 47<br>
slide44. Guidelines An application is submitted to TGA to register the drug in Australian Register of Therapeutic Goods (ARTG) after the completion of clinical trials.
The application consists of data to support the quality, safety and efficacy of the product for its intended use.
The application is assessed (on an administrative level) to make sure for compliance with basic guidelines and further evaluated by different sections and advice can also be sought on key issues to take final decision.
A company can make comments on the evaluation report, if necessary. 48<br>
slide45. 49 Preparation & submission of Application to DSEB of TGA Application reviewed by entry review team Accepted
Evaluation of Application -ve Sent back to
Applicant Within 225 days Report sent to TGA delegates Delegates seek advice of ADEC Delegates make final decision noting any recommendation from ADEC Approval of Drug Sent to Applicant Applicant comments on the report and submits supplementary data Applicant has an opportunity to
comment on delegates report Drug Registration Process in Australia<br>
slide46. COUNTRY SPECIFIC GUIDELINES 50<br>
slide47. Regulatory Agency State Food and Drug Administration (SFDA) Provincial Drug Administration Authorities (PDAAs)
▫ Organize the works of the formal review of submitted materials
Centre for Drug Evaluation (CDE)
▫ Evaluate the submitted information 51<br>
slide48. Drug Approval Process After formal review, the PDAAs send the qualified applications to the SFDA for further review.
The import drug registration application should be directly submitted to SFDA by the applicant.
SFDA’s Department of Drug Registration carefully reviews the completeness of the submitted materials, files the qualified applications and transmits all the materials of qualified applications to the directly attached to SFDA.
CDE determine whether the safety and effectiveness information submitted for a new drug are adequate for manufacturing and marketing approval and send the report of review to SFDA.
SFDA carefully considers the recommendations of CDE and makes a decision whether or not the drug registration application can be approved and issues the certificate of drug approval and drug approval number to the qualified applicant. 52<br>
slide49. 53 Dossier transferred from CDE to SFDA for administrative approval Regulatory approval granted Within 30 days Resubmission of amended application by applicant Technical review report of amended application by CDE Opinion of CDE on technical review Filing drug registration application to PDAA
After formal review
Application sent to SFDA for reviewing the completeness
Within 30 working days
Dossier transferred from SFDA to CDE for technical review
If complete within 120 days If not complete
Notify applicant for deficiency Within 4 months Within 40 days +ve -ve
Refusal to Proceed New Drug Registration Process of China<br>
slide50. COUNTRY SPECIFIC GUIDELINES 54<br>
slide51. Markets Brazil, Mexico, Venezuela , Ecuador and Colombia are some of the major pharmaceutical markets
Brazil and Mexico considered Regulated Markets
ANVISA : Stringent norms comparable to USFDA
Brazil : As per ANVISAs norms, CROs used for BE studies have to be approved and certified by the Brazilian authority.
For BE studies, innovator product manufactured in Brazil has to be used for comparison
ANVISA has implemented generic law to facilitate generic
market entry into the country
Mexico: Extensive analytical raw data of 3 stability batches.
Venezuela : Copies of source data 55<br>
slide52. Drug Regulatory Requirements - Technical Manufacturing Formula and Justification
Manufacturing Process
Process Validation
Batch Numbering
Batch Records
Stability Studies (Source Data)
▫ Accelerated
▫ Long Term 56<br>
slide53. Drug Regulatory Requirements - Legal Free Sale Certificate / Product Permission
GMP Certificate
Manufacturing License
Power of Attorney / Contract
Trademark Registration
Contract with local quality control facility 57<br>
slide54. Critical Success Factors US FDA / UK MCA / TGA approved facilities
Strong Regulatory Support
Good Distribution Set up
Price Competitiveness
Good Marketing Support
Strong group for registering products in place
Expansion in new markets and new products under feasibility studies 58<br>
slide55. Stability Issues Common Technical Document (CTD) norms are taken as
guidance for regulatory aspects during drug development
Major discrepancies in requirements of storage conditions for various countries
Stability studies on three batches - as per ICH guidelines / individual country norms
▫ Brazil: 30 ± 2 C; 75% ± 5% RH
▫ Colombia, Peru, Ecuador:30 ± 2C / 65% ± 5% RH
▫ Others: 25 ± 2 C; 60% ± 5% RH 59<br>
slide56. Data requirements 60 Site Registration Yes Plant GMP approval Major countries do audit. (Brazil, Mexico, Colombia) Stability Zone Zone II & Zone IVb Stability guidelines reference ANVISA and ICH Stability data 6– 12 months BE study for generics Brazil: Against Brazil reference drug in any CRO approved by ANVISA. PE to be done in Brazil Mexico: Against Mexican reference, in Mexico Only. Others: The BE for Brazil /Mexico is normally accepted. Major Holdup Certificate of P’ceutical Pdt., Legalizations, Translations, GMP audits, local requirements, time delay Dossier format Country specific<br>
slide57. Time and cost for registering a drug 61<br>
slide58. COUNTRY SPECIFIC GUIDELINES 62<br>
slide59. Guidelines Russia has its own, unique regulatory system which does not correspond to U.S. or EU practices.
Cultural and language barriers often become a challenge to foreign companies attempting to register pharmaceuticals by themselves
Registration procedure is also quite complicated and the documents tend to be modified due to constant changes in the regulatory requirements.
Before exporting a pharmaceutical product to Russia it has to be registered at the Ministry of Health of the Russian Federation; Department for registration of pharmaceuticals of the Federal Service for Health Control 63<br>
slide60. Registration Procedure The best way to accomplish registration of a pharmaceutical product in Russia is trough one of the following types of companies:
▫ Company incorporated in Russia and belonging to the Indian parent company
▫ Russian distributor/authorized agent
▫ Consulting company 64<br>
slide61. Documentation For the registration procedure a company has to submit the following:
▫ General documents
▫ Certificates
▫ Information and test reports
▫ Samples and package 65<br>
slide62. General documents Application for the State registration of a pharmaceutical which includes the name of the pharmaceutical preparation, the name and the contact information of the manufacturer
Name of the pharmaceutical preparation, including international non-proprietary name, scientific name in Latin, trade name and main synonyms
List of active ingredients and components
Recommended dosage, instruction for use
Description of the drug and its packaging, shelf life and
storage conditions
Power of Attorney issued by the manufacturer to the authorized company for carrying out registration procedure (notarized original with apostil) 66<br>
slide63. Certificates Copy of Free Sales Certificate (must be notarized and apostilled)
Copy of the license of pharmaceutical manufacture (must be notarized and apostilled)
Copy of GMP certificate (must be notarized and apostilled)
Copy of Certificate of manufacturer registration in their own country (must be notarized and apostilled)
Original of Certificate of analysis of the drug and its active substance (must
be signed and stamped by manufacturer)
Copy of Certificate of trade mark (must be signed and stamped by the manufacturer)
Information of registration of the drug in the country of the manufacture
and in other countries
A certificate of analysis of the drug in the country of origin of the manufacturer, GMP certificate and information on registration of the drug in the foreign country. 67<br>
slide64. Information and test reports Summary of the method of the drug manufacturing (must be signed and stamped by manufacturer)
Complete description of the quantitative and qualitative control methods with references to the pharmacopoeia and specification (must be signed and stamped by manufacturer)
Stability data of three drug series – by date
Patterns of the spectrums and chromatograms of the drug
Report of the pharmacological (specific) activity study substantiated the indications for use which are formed and described in the instruction
Test report of the drug toxicity (acute, sub acute, sub chronic, chronic toxicity) 68<br>
slide65. Information and test reports Test report of the specific influences (cancerogenity, mutagenic and teratogenic effects, embryotoxicity, allergic and local-irritative effects)
Clinical trial report of the medicine usage in clinic (the information which concerns only the drug that is produced by this manufacturer)
Copies of publications of the medicine usage in clinics after its registration in the country of origin (the information which concerns only the drug that is produced by this manufacturer)
Report of pharmacokinetics of the pharmaceutical study and its
bioequivalence to the original drug
Summary information of the side effects, in comparison with other analogous medicines, used at the same indications
Instruction for use (must be signed and stamped by the manufacturer) 69<br>
slide66. Samples and package Information of the material used for package: Certificates of the packaging materials (must be signed and stamped by the manufacturer)
Colour design of internal and external packages (Original and Russian version)
Standard samples of the active substance for quality control
Standard and referenced samples of the drug for the binding examination of quality (should be in the standard package) 70<br>
slide67. Approval Procedure The Ministry of Health determines whether these approvals are sufficient for an exemption of the drug from clinical and other testing in Russia before issuing a registration certificate.
Officially Russia does not recognize FDA and EU certificates.
According to the Law on Medicines, a fee in form of a state duty is to be paid for state registration and the registration should take no longer than six month. 71<br>